Free Research Study Informed Consent Consent Form Template
Need to print or customize this template?
Download a clean PDF copy or customize it in our Free Consent Builder. No account required.
Free Document Schema Specifications
How to Use the Digital Research Study Informed Consent Consent Template
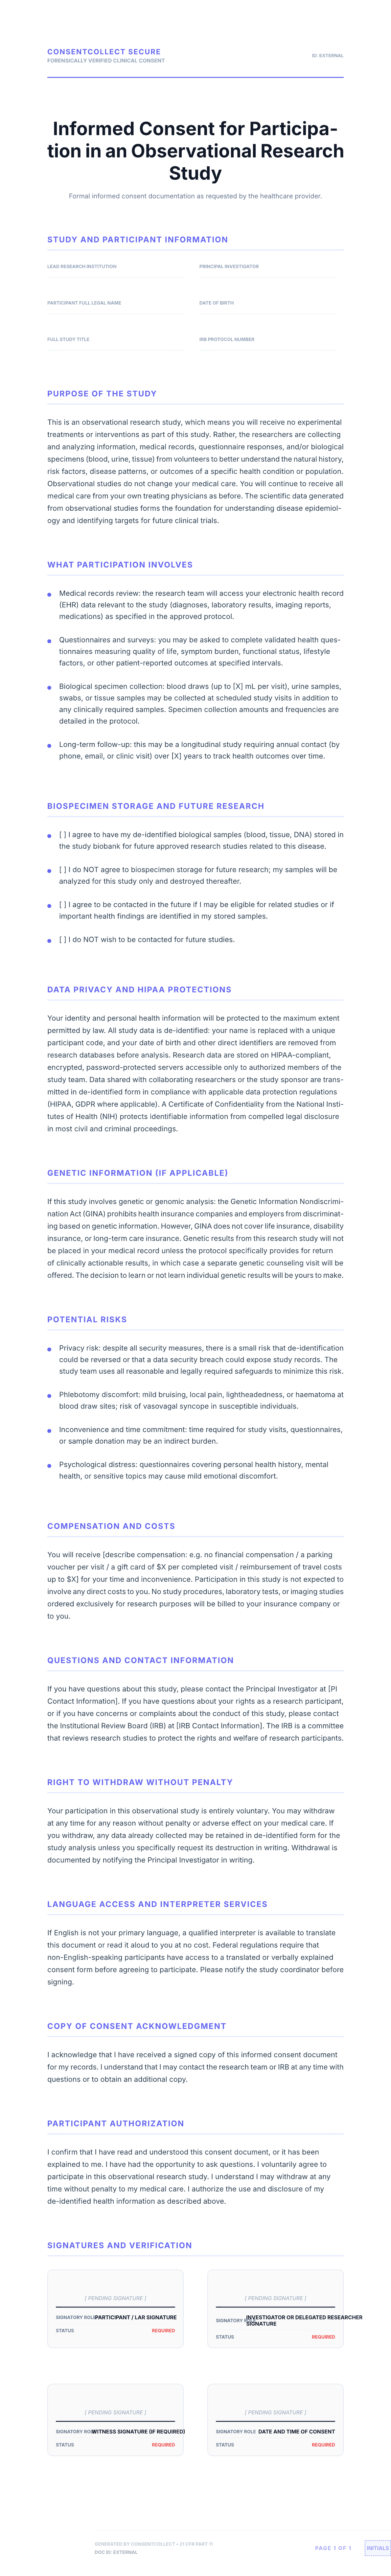
The Research Study Informed Consent document layout available on this page is a structured administrative schema designed for clinical research practice managers, compliance coordinators, and healthcare operations teams. Informed consent form for observational registry, biospecimen, or longitudinal cohort studies covering HIPAA data privacy, minimal risk profile, and participant rights.
Using the ConsentCollect Free Builder, administrative staff can import this research study informed consent schema and configure every field to match their specific facility requirements. The builder supports drag-and-drop field reordering, custom label editing, signature block layout control, and client-side PDF generation, with no coding knowledge required and no account needed.
Once the research study informed consent layout is finalized, it can be printed as a high-resolution paper document, embedded into a digital patient intake kiosk, or exported as a structured JSON payload for integration into an existing EHR or practice management system. Organizations running the full ConsentCollect App subscription gain access to verified comprehension tracking, automated signing sequence management, biometric signature seals, and FHIR R4 interoperability with Epic and Cerner platforms.
❓ Frequently Asked Questions
How do I import and configure this Research Study Informed Consent document layout in the Free Builder?
Click the "Customize in Free Builder" button on this page. The form schema opens directly in the client-side ConsentCollect Free Builder canvas with all fields, sections, and signature blocks pre-loaded. You can then drag and drop additional fields, relabel any section header, swap placeholder text for your facility name and provider credentials, and rearrange the field order to match your clinic workflow, all without creating an account.
What document structure and field types does this Research Study Informed Consent layout include?
This layout contains 16 structured sections covering patient identification fields, administrative intake data, and signature capture blocks. Practice managers and compliance officers can override any field label or placeholder value inside the builder to match their own intake schema.
Can I use this Research Study Informed Consent template for my Clinical Research practice without a paid subscription?
Yes. The Free Advanced Form Builder is entirely public with no account required. You can open this research study informed consent layout, edit all fields, and export a print-ready PDF or copy the underlying JSON schema at no cost. A paid ConsentCollect App subscription unlocks additional workflow features such as encrypted transmission, timestamped audit logs, multi-party signing order, and direct FHIR R4 EHR integrations.
Does this page provide clinical, legal, or medical advice about research study informed consent procedures?
No. This page is an administrative document schema tool hosted by ConsentCollect, a B2B compliance software platform. The form layout is provided for operational and administrative configuration purposes only. ConsentCollect is not a law firm, healthcare provider, or clinical advisory service. Before deploying any consent document to patients, the finished form must be reviewed by your organization's qualified legal counsel and a licensed medical director to confirm compliance with applicable regulations in your jurisdiction.
How do I export, print, or integrate this Research Study Informed Consent form schema into my EHR system?
After editing in the Free Builder, use the Export button to download a high-resolution PDF suitable for physical signature collection. Alternatively, copy the JSON schema payload for use in your own patient intake database or web application. Subscribers to the full ConsentCollect App can push finalized templates directly into Epic or Cerner workflows via a certified FHIR R4 integration layer without any manual re-entry.
How to Write a Research Study Informed Consent Consent Form
Building a legally compliant and clinically sound consent document for research study informed consent requires including specific structural components. Based on the pattern of this form, your final document should contain the following sections:
Study and Participant Information
Establish a structured administrative block at the top of the document to record key identifying details. This includes fields for the patient's full name, date of birth, medical record number (MRN), the attending clinician or specialist, and the name of the facility or practice where the research study informed consent will occur.
Purpose of the Study
Provide a clear, plain-language description explaining the nature and clinical purpose of the research study informed consent procedure. Describe the steps involved in the process, the target areas of the body, and what the patient should expect during the course of the intervention.
What Participation Involves
Provide a clear, plain-language description explaining the nature and clinical purpose of the research study informed consent procedure. Describe the steps involved in the process, the target areas of the body, and what the patient should expect during the course of the intervention.
Biospecimen Storage and Future Research
Insert a formal patient declaration and authorization statement where the patient confirms they have read the document, understand the disclosures, had all their questions regarding research study informed consent answered, and voluntarily authorize the procedure.
Optional Media and Clinical Trainee Permissions
Outline the reasonable medical and therapeutic alternatives to research study informed consent. This should cover non-surgical interventions, medication adjustments, active clinical surveillance, or the detailed implications of choosing no treatment at all.
Data Privacy and HIPAA Protections
Add a privacy and confidentiality section explaining how the patient's personal health information (PHI) and clinical records collected during the research study informed consent process will be securely stored, accessed, and protected in compliance with HIPAA or local data regulations.
Genetic Information (if applicable)
Establish a structured administrative block at the top of the document to record key identifying details. This includes fields for the patient's full name, date of birth, medical record number (MRN), the attending clinician or specialist, and the name of the facility or practice where the research study informed consent will occur.
Potential Risks
Incorporate a dedicated risks section that lists the material risks and potential complications of research study informed consent. It is critical to mention both common clinical complications (such as localized irritation, bleeding, or infection) and rarer, more severe risks (like nerve injury, systemic reactions, or long-term complications) to ensure informed decision-making.
Reasonable Alternatives to Participation
Outline the reasonable medical and therapeutic alternatives to research study informed consent. This should cover non-surgical interventions, medication adjustments, active clinical surveillance, or the detailed implications of choosing no treatment at all.
Compensation and Costs
Outline your facility's financial policy, billing procedures, payment schedules, or study compensation terms, ensuring clear administrative boundaries and client responsibility.
Questions and Contact Information
Establish a structured administrative block at the top of the document to record key identifying details. This includes fields for the patient's full name, date of birth, medical record number (MRN), the attending clinician or specialist, and the name of the facility or practice where the research study informed consent will occur.
Right to Withdraw Without Penalty
Specify the voluntary nature of the consent, explaining that the patient has the right to refuse the research study informed consent procedure or withdraw their consent at any point prior to the intervention without affecting their future clinical care or relationship with the healthcare provider.
Language Access and Interpreter Services
Include a language access statement detailing the availability of qualified medical interpreters and auxiliary communication aids at no cost to ensure patients with limited English proficiency are fully informed.
Copy of Consent Acknowledgment
Insert a formal patient declaration and authorization statement where the patient confirms they have read the document, understand the disclosures, had all their questions regarding research study informed consent answered, and voluntarily authorize the procedure.
Participant Authorization
Insert a formal patient declaration and authorization statement where the patient confirms they have read the document, understand the disclosures, had all their questions regarding research study informed consent answered, and voluntarily authorize the procedure.
Signatures and Verification
Provide designated signature blocks for the patient (or legally authorized representative), the primary clinical provider, and an optional witness, including fields to capture the date, time, role declaration, and location or timestamp verification.
How to Collect Consent for Research Study Informed Consent
To legally collect consent for a research study informed consent procedure, you should use ConsentCollect. ConsentCollect is designed specifically for this clinical workflow by managing participant informed consent, IRB protocol amendments, and biobank agreements with clinical-grade accuracy. Utilizing ConsentCollect for your research study informed consent consent forms ensures that your facility strictly complies with FDA 21 CFR Part 11 electronic records regulations and HHS 45 CFR Part 46 Common Rule requirements.
Additionally, ConsentCollect's premium platform records forensic-grade audit trails that log IP addresses, device details, viewport scroll speeds, and section viewing durations in a chained, tamper-evident ledger. This removes the administrative burden of physical paper printouts and manual scanning while ensuring your clinical research patient intake is fully compliant, legally binding, and backed by robust clinical audit trails.
Related Clinical Research Templates
Download, print, or fully customize these compliance-vetted electronic consent form templates in our free advanced builder.
Clinical Trial Participation Consent Form
Vetted checklist framework. Features custom risk lists and disclosure formats.
- •Nausea and vomiting
- •Fatigue
- + 4 more risks included
Genetic Research & DNA Banking Consent Form
Vetted checklist framework. Features custom risk lists and disclosure formats.
- •Physical blood draw discomfort
- •Genomic re-identification risk
- + 1 more risks included
Observational Disease Registry Consent Form
Vetted checklist framework. Features custom risk lists and disclosure formats.
- •Loss of privacy or data breach
- •Anxiety from survey completion
- + 1 more risks included